Category Archives: Molecular dynamics simulatons

First-Principles Simulation of Ionizing Radiation Damage in Molecular Systems over Short Time Scales

Naija Pison, Aurelio Alvarez-Ibarra, Roberto Flores-Moreno, Dominique Guillaumont, Aurélien de la Lande, J. Chem. Theor. Comput. 2026 DOI: doi.org/10.1021/acs.jctc.6c01190

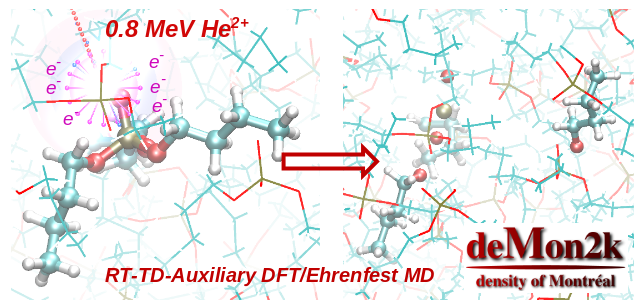
In this work, we present an ab initio molecular simulation method designed to investigate the ultrashort timescales of radiation chemistry induced by ionizing radiation, in particular fast ions and electric pulses. The approach accounts for energy deposition at the sub-femtosecond scale, electron emission, and energy dissipation into nuclear vibrational modes, which may ultimately lead to the formation of the earliest radiation-induced damages. The method, implemented in deMon2k, is based on the coupling between Real-Time Time-Dependent Density Functional Theory (RT-TDDFT) within the Auxiliary Density Functional Theory formalism and Ehrenfest mean-field dynamics. A hybrid QM/MM scheme further enables the simulation of radiation damage formation in complex and heterogeneous systems such as proteins or DNA. The method has been tested on a set of benchmark molecules and achieves controlled numerical accuracy down to the micromicroeV scale for the total energy conservation. From a computational efficiency perspective, the implementation relies on the Auxiliary DFT formalism and fully benefits from modern CPU- and GPU-based computer architectures. As an illustrative application, we present simulations of a solvated tributyl phosphate molecule subjected to alpha radiation, a problem of direct relevance to the nuclear industry. The study analyzes the energy deposition mechanisms, the dissipation pathways, and the characterization of the earliest radiation-induced damages. In summary, the ionizing-radiation chemistry module of deMon2k presented here opens the way toward a deeper understanding of the ultrafast formation of radiation-induced chemical damages in fields as diverse as biology, the nuclear industry, aerospace science, and astrochemistry.
AMOEBA Polarizable Atomic Multipole Force Field Parameters for Flavins

Mélody Aeck, Aurélien de la Lande, Carine Clavaguéra, Fabien Cailliez. J. Chem. Theory Comput. 2026 22, 8954–8965. DOI: https://doi.org/10.1021/acs.jctc.6c00958.

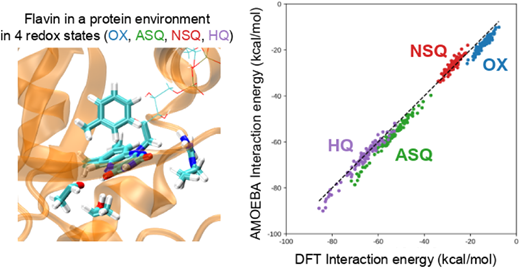
Computation of redox potential of cofactors in biological systems requires an accurate description of the interaction between the redox cofactors and the biomolecular environment and extensive conformational sampling. Seeking to achieve a quantitative leap in redox potential evaluation for flavoproteins, we report here the calibration of sets of parameters to describe flavin molecules in various redox states in the framework of AMOEBA, a multipolar and polarizable force field. We calibrate both the isoalloxazine ring in the five biologically relevant redox states and the polyol chain needed to describe riboflavin or flavin adenine dinucleotide molecules. We propose a delineated, step-by-step protocol to accurately account for the physical effects at play. We provide evidence that these parameters produce stable conformations in agreement with quantum chemistry (DFT) calculations. The newly derived AMOEBA force field allows to compute interaction energies of lumiflavin with water molecules and amino acids in very good agreement with quantum chemistry calculations, with a net improvement with respect to a monopolar nonpolarizable force field. Finally, as a first application, we compute redox free energy of half-reactions for two redox couples of lumiflavin in water. The values obtained are once again in better agreement with DFT values than those obtained with a standard classical force field.

Computational Study of Heme b595 to Heme d Electron Transfer in E. coli Cytochrome bd-I Oxidase

Raaif Siddeeque, Baptiste Etcheverry, Côme Cattin, Jean Deviers, Frédéric Melin, Petra Hellwig, Fabien Cailliez, Aurélien de la Lande. J. Chem. Inform. Mod. 2026, 66, 1757. DOI: doi.org/10.1021/acs.jcim.5c02194. Link to Biorxiv

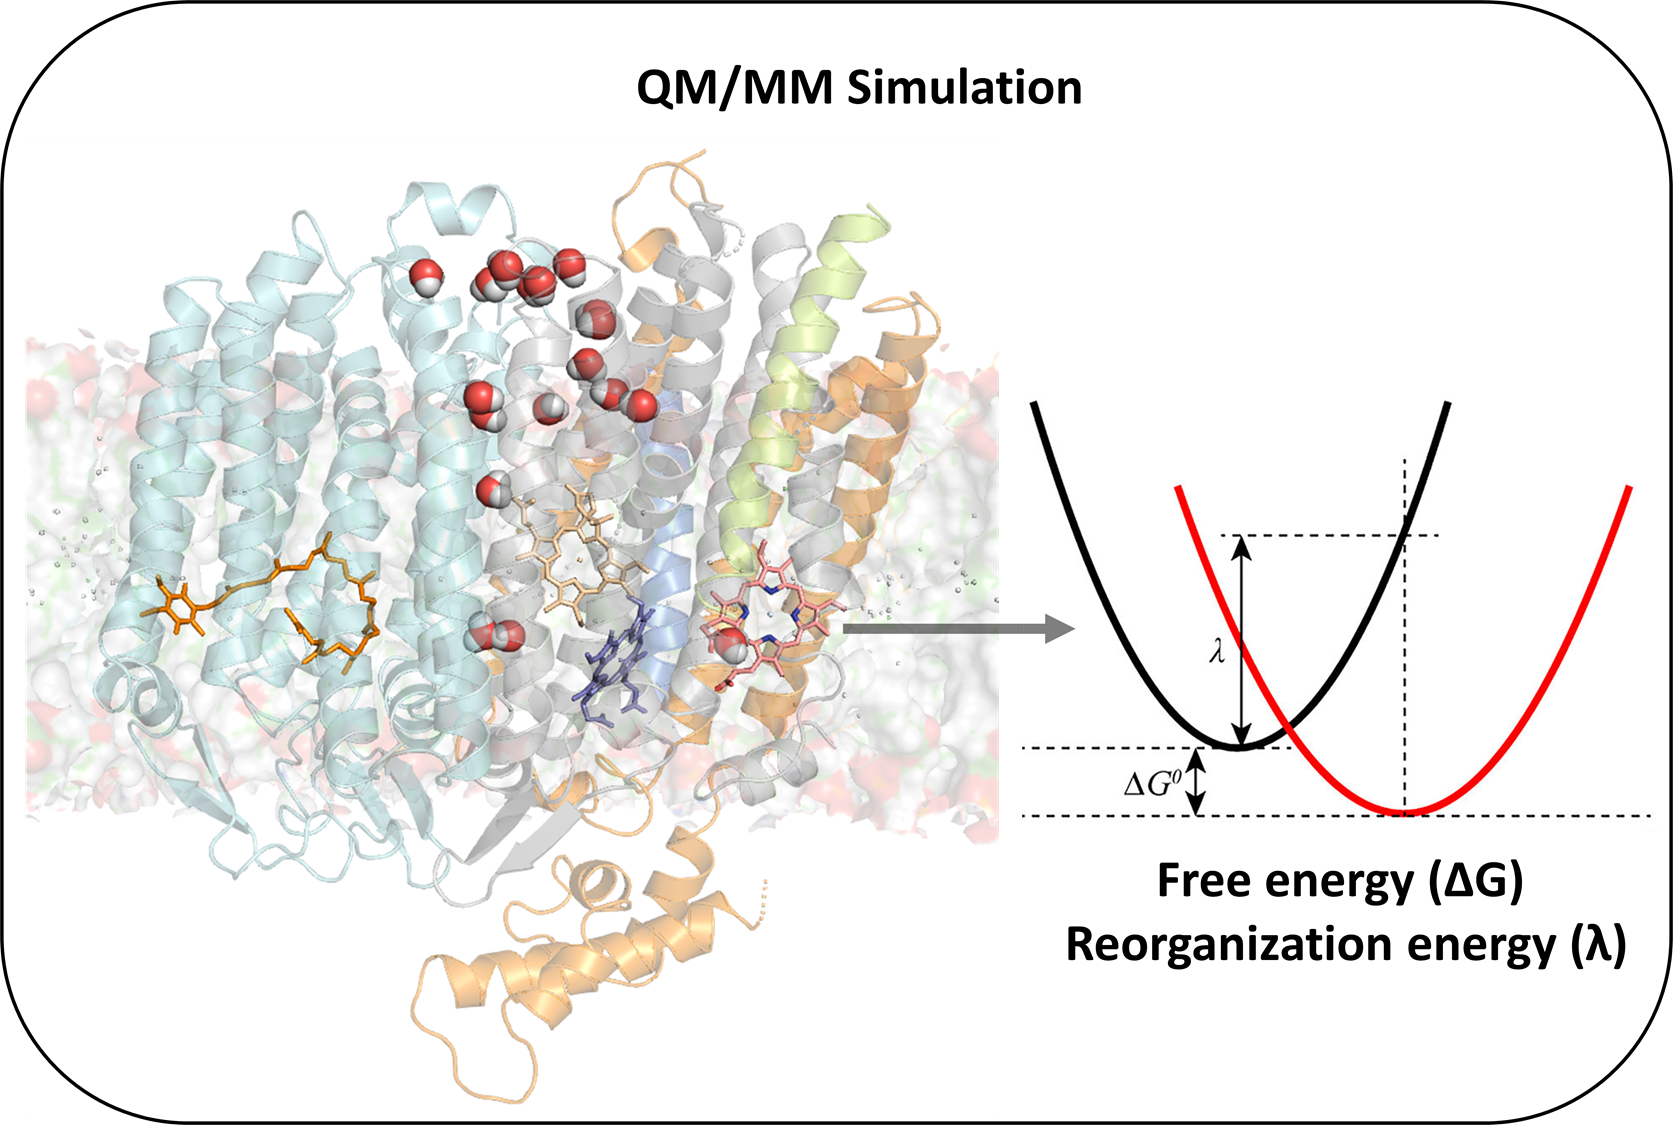
Cytochrome bd is a distinctive family of terminal oxidases present in the respiratory chains of many prokaryotes. Despite their biological importance, the redox chemistry of these proteins remains poorly understood, largely due to the presence of two b-type hemes and one d-type heme. Here, we report the first computational study of interheme electron transfer in the cytochrome bd family. We performed 10 μs of molecular dynamics simulations of E. coli cytochrome bd-I embedded in realistic membranes, combined with quantum chemical calculations to estimate the thermodynamic parameters of electron transfer from heme b595 to heme d within the framework of Marcus theory. We further identify the respective contributions of the hemes, protein scaffold, lipid bilayer, water, and counterions to the driving force and reorganization energy. The interheme electronic coupling was calculated using the Projected Orbital Diabatization (POD) method in a hybrid Quantum Mechanics/Molecular Mechanics scheme and rationalized through electron transfer pathway analysis. This study provides fundamental insights into how electron transfer steps are orchestrated in the catalytic cycle of E. coli cytochrome bd-I.

NOX transmembrane electron transfer is governed by a subtly balanced, self-adjusting charge distribution

Baptiste Etcheverry, Marc baaden, Aurélien de la Lande, Fabien Cailliez, under review. Link to BioXxiv

NADPH oxidases (NOX) form a family of transmembrane enzymes that catalyze the formation of reactive oxygen species. These are produced thanks to a chain of electron transfers (ET), shuttling electrons from one side of the membrane to the other, using one flavin and two heme cofactors as redox mediators. In this work we investigate the thermodynamics of the electron transfer (ET) between the two hemes contained in the transmembrane domain by means of extensive molecular dynamics simulations. We compare two proteins of the NOX5 isoform, from homo sapiens (hNOX5) and from cylindrospermum stagnale (csNOX5), a cyanobacteria. We study in detail the influence of both the density of negatively charged lipids in the membrane and of the NOX5 aminoacid sequence on the ET thermodynamic balance. The linear response formalism allows us to decompose the variation in free energy into the individual contributions of the system components (protein, membrane, solvent, etc.). We highlight the major compensatory effects of the various components in the global free energy budget in those complex systems. Although the contributions of the protein or the membrane to the ET thermodynamics can be individually strongly modified by a change in the aminoacid sequence or the membrane composition, they are largely compensated by the rest of the heme environment so that the total free energy is always found to be slightly favorable to the electron transfer. To our knowledge, this study is the first to highlight the effect of membrane charge density on inter-heme ET, providing valuable insights into the molecular mechanisms governing ET catalysis in complex membrane systems.

Avian cryptochrome 4 binds superoxide

Jean Deviers, Fabien Cailliez, Aurélien de la Lande, Daniel R Kattnig, Comput. Struct. Biol. J. 2024, 26, 11-21. https://doi.org/10.1016/j.csbj.2023.12.009 (open access).

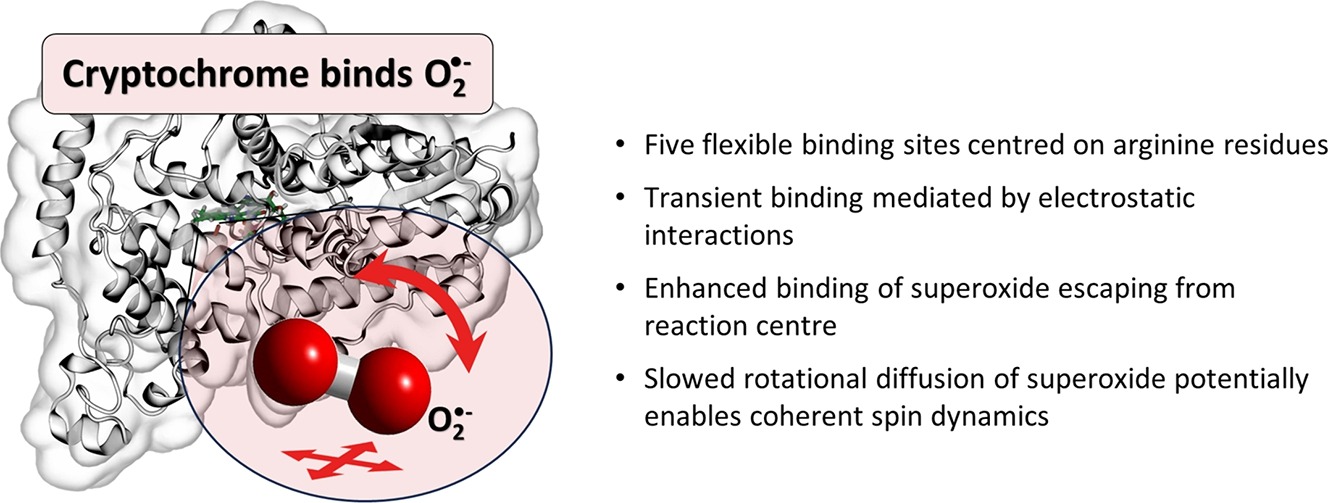
Flavin-binding cryptochromes are blue-light sensitive photoreceptors that have been implicated with magnetoreception in some species. The photocycle involves an intra-protein photo-reduction of the flavin cofactor, generating a magnetosensitive radical pair, and its subsequent re-oxidation. Superoxide (O) is generated in the re-oxidation with molecular oxygen. The resulting O-containing radical pairs have also been hypothesised to underpin various magnetosensitive traits, but due to fast spin relaxation when tumbling in solution would require immobilisation. We here describe our insights in the binding of superoxide to cryptochrome 4 from C. livia based on extensive all-atom molecular dynamics studies and density-functional theory calculations. The positively charged “crypt” region that leads to the flavin binding pocket transiently binds O at 5 flexible binding sites centred on arginine residues. Typical binding times amounted to tens of nanoseconds, but exceptional binding events extended to several hundreds of nanoseconds and slowed the rotational diffusion, thereby realising rotational correlation times as large as 1 ns. The binding sites are particularly efficient in scavenging superoxide escaping from a putative generation site close to the flavin-cofactor, possibly implying a functional relevance. We discuss our findings in view of a potential magnetosensitivity of biological flavin semiquinone/superoxide radical pairs.