Roberta Leonetti, Federica Agostini, Aurélien de la Lande, J. Chem. Phys. 165, 094119 (2026).
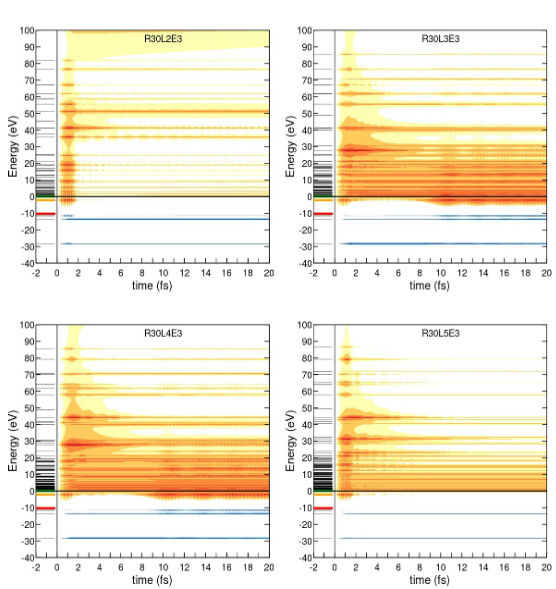
We explore the capability of real-time time-dependent auxiliary density functional theory (RT-TD-ADFT) to capture photoionization of the nitrogen molecule when employed in combination with atomic Gaussian basis sets optimized for the continuum (AOC). The molecule is ionized by an ultrashort laser pulse in the near- and far off-resonant energy conditions. The dependence of the calculated amount of deposited energy and of emitted electrons on various sets of AOC is investigated. We find that for energies below a threshold of ∼25 eV, the simulations carried out with sufficiently angularly flexible AOC capture reasonably well the physics of the photoionization process, while AOC do not seem adequate to describe photoionization occurring at higher energies. We also find that accounting for exact exchange in the simulation moderately affects the results in the near-resonant situation, while it has a stronger impact on the appearance of the so-called molecular dipole instability a few fs after ionization in the far off-resonant case. Overall, this work paves the way for the evaluation of photoelectron spectra based on RT-TD-ADFT.